Sentieon
Sentieon 中文手册
Sentieon 中文手册(上册)
Sentieon 中文手册(下册)
Sentieon 软件应用教程
Sentieon | 联合Dell、AMD开发基因组大数据分析加速方案
Sentieon | 应用教程: 利用Sentieon Python API引擎为自研算法加速
Sentieon | 应用教程: 关于读段组的建议
Sentieon | 应用教程: 使用DNAscope对HiFi长读长数据进行胚系变异检测分析
Sentieon | 应用教程: TNscope® 使用机器学习模型进行有匹配正常样本的体细胞变异发现
Sentieon | 应用教程: CCDG使用Sentieon®的功能等效流程
Sentieon | 应用教程: 利用共识功能去除PCR重复
Sentieon | 应用教程: 适用于PacBio HiFi和Oxford Nanopore长读长测序数据的结构变异检测
Sentieon | 应用教程: 使用 Sentieon进行大型基因组重测序分析
Sentieon | 应用教程: 体细胞SNP/Indel变异检测
Sentieon | 应用教程: DNAscope使用机器学习模型进行胚系变异调用
Sentieon | 应用教程: 唯一分子标识符(UMI)
Sentieon | 应用教程: Sentieon分布模式
Sentieon | 应用教程:使用CNVscope进行CNV检测分析
Sentieon发布核心家系(trio)基因分析最佳实践方案
Sentieon推出Segdup-caller:针对片段重复区域的专用精准变异检测工具
Sentieon-cli | DNAscope WES 流程单条命令版本详解
Sentieon | DNAscope 核心家系(trio) WES 分析全流程详解
Sentieon | DNAscope Hybrid长短读长混合分析流程详解及评测
Sentieon | DNAscope LongRead:面向ONT WGS的快速、高精度、低成本变异检测流程
重磅发布|真迈生物联合Sentieon打造SURFSeq 5000 CMS + DNAscope工作流
Sentieon软件版本更新
Sentieon | 发布V202503.01版本
Sentieon | 发布V202503.02版本
Sentieon | 发布V202503.03版本
Sentieon | 发布V202503.04版本
Sentieon软件快速入门指南
Sentieon 软件模块总述
Sentieon 特色流程 - DNAscope
Sentieon | DNAscope Illumina 流程
sentieon | DNAscope Complete Genomics 流程
Sentieon | DNAscope LongRead PacBio 流程
Sentieon | DNAscope Ultima Genomics 流程
Sentieon | DNAscope Element Bio 流程
Sentieon | DNAscope LongRead Nanopore 流程
Sentieon混合分析流程 - DNAscope Hybrid
Sentieon推出混合型短读长和长读长变异检测DNAscope Hybrid流程(上)
Sentieon推出混合型短读长和长读长变异检测DNAscope Hybrid流程(下)
毅硕Sentieon | 泛基因组分析流程详解
Sentieon | 基于Illumina WGS数据的高精度、低成本的泛基因组分析方案
Sentieon | 基于Ultima WGS数据的高精度、低成本的泛基因组分析测评
Sentieon | 泛基因组分析流程详解—CHM13 (T2T) 更新版
Sentieon | DNAscope Pangenome:赋能人类高精度生殖筛查
Sentieon|植物泛基因组分析加速详解
毅硕Sentieon | RNA-seq 变异检测全流程详解
毅硕Sentieon | 物种全基因组(WGS)分析流程
毅硕Sentieon | 植物全基因组(WGS)分析流程
毅硕Sentieon | 小麦(Triticum aestivum)全基因组WGS分析流程
毅硕Sentieon | 水稻(Oryza sativa)全基因组WGS分析流程
毅硕Sentieon | 拟南芥(Arabidopsis thaliana)全基因组WGS分析流程
毅硕Sentieon | 马铃薯(Solanum tuberosum)全基因组WGS分析流程
毅硕Sentieon | 巨桉(Eucalyptus grandis)全基因组WGS分析流程
毅硕Sentieon | 向日葵(Helianthus annuus)全基因组WGS分析流程
毅硕Sentieon | 野草莓(Fragaria vesca)全基因组WGS分析流程
毅硕Sentieon | 银杏(Ginkgo biloba)全基因组WGS分析流程
毅硕Sentieon | 大豆(Glycine max)全基因组WGS分析流程
毅硕Sentieon | 陆地棉(Gossypium hirsutum)全基因组WGS分析流程
毅硕Sentieon | 胡桃(Juglans regia)全基因组WGS分析流程
毅硕Sentieon | 苹果(Malus domestica)全基因组WGS分析流程
毅硕Sentieon | 香蕉(musa acuminata)全基因组WGS分析流程
毅硕Sentieon | 罂粟(Papaver somniferum)全基因组WGS分析流程
毅硕Sentieon | 开心果(Pistacia vera)全基因组WGS分析流程
毅硕Sentieon | 毛果杨(Populus trichocarpa)全基因组WGS分析流程
毅硕Sentieon | 夏栎(Quercus robur)全基因组WGS分析流程
毅硕Sentieon | 蓖麻(Ricinus communis)全基因组WGS分析流程
毅硕Sentieon | 动物全基因组(WGS)分析流程
毅硕Sentieon | 猪(sus scrofa)全基因组WGS分析流程
毅硕Sentieon | 鸡(Gallus gallus)全基因组WGS分析流程
毅硕Sentieon | 家鼠(Mus musculus)全基因组WGS分析流程
毅硕Sentieon | 家犬(canis lupus familiaris)全基因组WGS分析流程
毅硕Sentieon | 东方蜜蜂(Apis cerana)全基因组WGS分析流程
毅硕Sentieon | 电鳗(Electrophorus electricus)全基因组WGS分析流程
毅硕Sentieon | 红隼(Falco tinnunculus)全基因组WGS分析流程
毅硕Sentieon | 家猫(Felis catus)全基因组WGS分析流程
毅硕Sentieon | 尼罗罗非鱼(Oreochromis niloticus)全基因组WGS分析流程
毅硕Sentieon | 狮子(Panthera leo)全基因组WGS分析流程
毅硕Sentieon | 东北虎(panthera tigris altaica)全基因组WGS分析流程
毅硕Sentieon | 黑猩猩(pan troglodytes)全基因组WGS分析流程
毅硕Sentieon | 中国鳖(pelodiscus sinensis)全基因组WGS分析流程
毅硕Sentieon | 褐家鼠(rattus norvegicus)全基因组WGS分析流程
毅硕Sentieon | 家牛(Bos taurus)全基因WGS分析流程
毅硕Sentieon | 虎河豚(takifugu rubripes)全基因WGS分析流程
毅硕Sentieon | 人类(Homo sapiens)全基因组WGS分析流程
毅硕Sentieon | 狮尾狒(theropithecus gelada)全基因组WGS分析流程
毅硕Sentieon | 草鱼(Ctenopharyngodon idella)全基因组WGS分析流程
毅硕Sentieon文献解读
Sentieon文献解读 | Population Sequencing
Sentieon文献解读 | Agrigenomics
Sentieon | Agrigenomics-泛基因组揭示小麦结构变异与栖息地及育种的关联
Sentieon文献解读 | Genetic Disease
Sentieon文献解读 | Tumor Sequencing
Sentieon文献解读 | Benchmark and Method Study
Sentieon文献解读 | Long Read Sequencing
Sentieon文献解读 | Clinical Trial
Sentieon文献解读 | Epidemiology
Sentieon文献解读 | Gene Editing
Sentieon文献解读 | Liquid Biopsy
Sentieon | TNscope 分析流程详解
Sentieon | TNscope WES Panel 分析流程详解
-
+
home
Sentieon | TNscope 分析流程详解
# 一、前言 在肿瘤基因组学研究中,精准识别体细胞变异位点(call somatic mutations)是解析肿瘤发生机制、寻找用药靶点与监控疗效的核心基础。然而,由于肿瘤样本的异质性和测序噪音,高灵敏度和高特异性的变异检测极具挑战。 **Sentieon 的 TNscope 模块为此提供了高效解决方案,TNscope 流程采用了优化的机器学习算法,能在极大提升分析速度的同时,显著提高体细胞点突变与插入缺失检测的灵敏度与准确性,尤其擅长处理低丰度变异。该模块已成为肿瘤精准医疗与大规模测序项目中不可或缺的生信分析工具。** **下图展示了利用 Sentieon 软件的 TNscope 模块进行体细胞变异和结构变异检测,从而运行仅针对肿瘤或肿瘤-正常组织样本的生物信息学分析流程。** 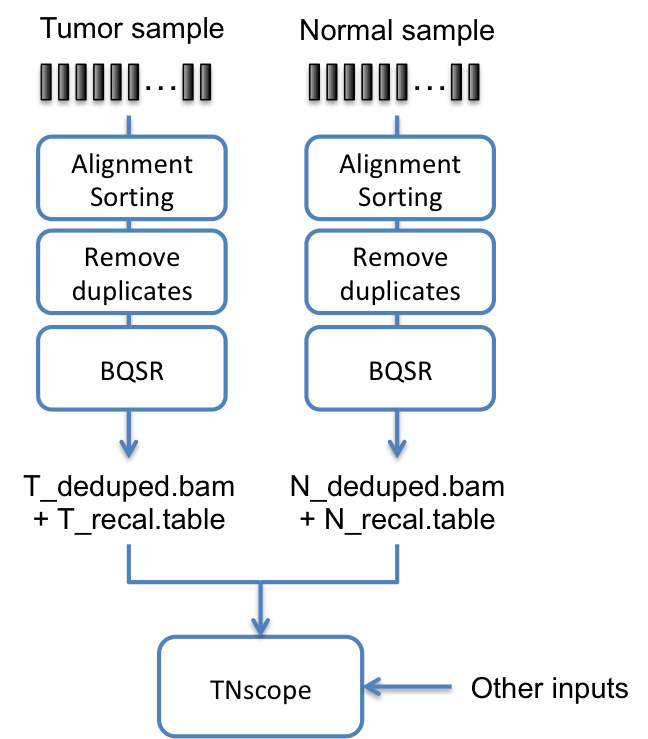 *图1 Sentieon TNscope 的生物信息学分析流程* *** # 二、流程总览 ## 1. 概述 在这个生物信息学流程中,你需要以下输入: - 参考基因组 FASTA 文件。 - 两组包含待分析样本核苷酸序列的 FASTQ 文件,一组用于肿瘤样本,一组用于匹配正常样本。这些文件包含 DNA 测序的原始读段。该软件支持输入使用 GZIP 压缩的 FASTQ 文件。该软件仅支持包含桑格格式(Phred+33)高质量评分的文件。 - (可选)单核苷酸多态性数据库(dbSNP)数据。数据以 VCF 文件的形式使用。 - (可选)多个已知位点数据。数据以 VCF 文件的形式使用。 以下步骤构成肿瘤正常匹配对的典型生物信息学流程: - 对肿瘤样本和正常样本,分别采用类似于“DNAseq® 典型使用流程”中所介绍的 DNA 测序流程,进行独立的前处理,包括以下步骤: - 将测序读数比对到参考基因组上。你需要确保肿瘤样本和正常样本的 SM 标签不同,因为在体细胞变异检测中,这将被用作一个参数。同时,还需要确保两个样本的 `RG:ID` 是不同且唯一的。 - 统计数据指标。 - 删除重复序列。 - (可选)基础质量评分重新校准(BQSR)。 - 体细胞变异检测:此步骤旨在识别肿瘤基因组数据中相较于正常基因组存在体细胞变异的位点,并计算该位点的基因型。 ## 2. 分析流程步骤 ### (1)比对 使用多线程时,为了保持 bwa 输出结果的一致性,需要添加 `-K 10000000` 参数。 ``` # ****************************************** # 1a. Mapping reads with BWA-MEM, sorting for tumor sample # ****************************************** # The results of this call are dependent on the number of threads used. # To have number of threads independent results, add chunk size option -K 10000000 ( $SENTIEON_INSTALL_DIR/bin/sentieon bwa mem \ -R "@RG\tID:$TUMOR_RGID\tSM:$TUMOR_SM\tPL:$PL" \ -t $NT -K 10000000 $FASTA $TUMOR_FASTQ_1 $TUMOR_FASTQ_2 || \ { echo -n 'BWA error'; exit 1; } ) | \ $SENTIEON_INSTALL_DIR/bin/sentieon util sort \ -o tumor_sorted.bam -t $NT --sam2bam -i - || \ { echo "Alignment1 failed"; exit 1; } # ****************************************** # 1b. Mapping reads with BWA-MEM, sorting for normal sample # ****************************************** # The results of this call are dependent on the number of threads used. # To have number of threads independent results, add chunk size option -K 10000000 ( $SENTIEON_INSTALL_DIR/bin/sentieon bwa mem \ -R "@RG\tID:$NORMAL_RGID\tSM:$NORMAL_SM\tPL:$PL" \ -t $NT -K 10000000 $FASTA $NORMAL_FASTQ_1 $NORMAL_FASTQ_2 || \ { echo -n 'BWA error'; exit 1; } ) | \ $SENTIEON_INSTALL_DIR/bin/sentieon util sort \ -o normal_sorted.bam -t $NT --sam2bam -i - || \ { echo "Alignment2 failed"; exit 1; } ``` ### (2)统计质控和质控绘图 ``` # ****************************************** # 2a. Metrics for tumor sample # ****************************************** $SENTIEON_INSTALL_DIR/bin/sentieon driver -r $FASTA -t $NT -i tumor_sorted.bam \ --algo MeanQualityByCycle tumor_mq_metrics.txt \ --algo QualDistribution tumor_qd_metrics.txt \ --algo GCBias --summary tumor_gc_summary.txt tumor_gc_metrics.txt \ --algo AlignmentStat --adapter_seq '' tumor_aln_metrics.txt \ --algo InsertSizeMetricAlgo tumor_is_metrics.txt \ --algo CoverageMetrics tumor_coverage_metrics || \ { echo "Metrics1 failed"; exit 1; } $SENTIEON_INSTALL_DIR/bin/sentieon plot GCBias \ -o tumor_gc-report.pdf tumor_gc_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot QualDistribution \ -o tumor_qd-report.pdf tumor_qd_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot MeanQualityByCycle \ -o tumor_mq-report.pdf tumor_mq_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot InsertSizeMetricAlgo \ -o tumor_is-report.pdf tumor_is_metrics.txt # ****************************************** # 2b. Metrics for normal sample # ****************************************** $SENTIEON_INSTALL_DIR/bin/sentieon driver -r $FASTA -t $NT -i normal_sorted.bam \ --algo MeanQualityByCycle normal_mq_metrics.txt \ --algo QualDistribution normal_qd_metrics.txt \ --algo GCBias --summary normal_gc_summary.txt normal_gc_metrics.txt \ --algo AlignmentStat --adapter_seq '' normal_aln_metrics.txt \ --algo InsertSizeMetricAlgo normal_is_metrics.txt \ --algo CoverageMetrics normal_coverage_metrics || \ { echo "Metrics2 failed"; exit 1; } $SENTIEON_INSTALL_DIR/bin/sentieon plot GCBias \ -o normal_gc-report.pdf normal_gc_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot QualDistribution \ -o normal_qd-report.pdf normal_qd_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot MeanQualityByCycle \ -o normal_mq-report.pdf normal_mq_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon plot InsertSizeMetricAlgo \ -o normal_is-report.pdf normal_is_metrics.txt ``` 如果输入样本是 WGS,也可以添加一步统计: ``` $SENTIEON_INSTALL_DIR/bin/sentieon driver --temp_dir $TMPDIR -r $FASTA -t $NT \ -i tumor_sorted.bam --algo WgsMetricsAlgo tumor.wgs_metrics.txt $SENTIEON_INSTALL_DIR/bin/sentieon driver --temp_dir $TMPDIR -r $FASTA -t $NT \ -i normal_sorted.bam --algo WgsMetricsAlgo normal.wgs_metrics.txt ``` ### (3)标记重复或去重复 Dedup 模块默认是标记重复 reads。如果需要删除重复 reads,可以添加 `--rmdup` 参数。 ``` # ****************************************** # 3a. Remove Duplicate Reads for tumor sample. # It is possible to remove instead of mark duplicates by adding the --rmdup option in Dedup. # ****************************************** $SENTIEON_INSTALL_DIR/bin/sentieon driver -t $NT -i tumor_sorted.bam \ --algo LocusCollector --fun score_info tumor_score.txt || \ { echo "LocusCollector1 failed"; exit 1; } $SENTIEON_INSTALL_DIR/bin/sentieon driver -t $NT -i tumor_sorted.bam \ --algo Dedup --score_info tumor_score.txt \ --metrics tumor_dedup_metrics.txt tumor_deduped.bam || \ { echo "Dedup1 failed"; exit 1; } # ****************************************** # 3b. Remove Duplicate Reads for normal sample. # It is possible to remove instead of mark duplicates by adding the --rmdup option in Dedup. # ****************************************** $SENTIEON_INSTALL_DIR/bin/sentieon driver -t $NT -i normal_sorted.bam \ --algo LocusCollector --fun score_info normal_score.txt || \ { echo "LocusCollector2 failed"; exit 1; } $SENTIEON_INSTALL_DIR/bin/sentieon driver -t $NT -i normal_sorted.bam \ --algo Dedup --score_info normal_score.txt \ --metrics normal_dedup_metrics.txt normal_deduped.bam || \ { echo "Dedup2 failed"; exit 1; } ``` ## 3. 兼顾配对与无配对样本的体细胞变异分析 只需执行一个指令,即可调用肿瘤-正常匹配对的变异。 ``` sentieon driver -t NUMBER_THREADS -r REFERENCE \ -i TUMOR_DEDUPED_BAM [-q TUMOR_RECAL_DATA.TABLE] \ -i NORMAL_DEDUPED_BAM [-q NORMAL_RECAL_DATA.TABLE] \ --algo TNscope \ --tumor_sample TUMOR_SAMPLE_NAME --normal_sample NORMAL_SAMPLE_NAME \ [--dbsnp DBSNP] OUT_TN_VCF ``` 如果没有匹配的正常采样,可以跳过 Normal BAM 文件和 Normal 名输入,因为这些不是必需的;但由于缺乏正常样本,生殖系变异会出现在输出文件中: ``` sentieon driver -t NUMBER_THREADS -r REFERENCE \ -i TUMOR_DEDUPED_BAM [-q TUMOR_RECAL_DATA.TABLE] \ --algo TNscope --tumor_sample TUMOR_SAMPLE_NAME \ [--dbsnp DBSNP] OUT_TN_VCF ``` 为了在缺少匹配正常样本时过滤生殖系变异,可以用通用 PON 替换匹配的正常样本,使用下文所述方法生成 PON VCF(Panel of Normal VCF)文件。 只需执行一个命令即可检测 tumor only 样本上的变异: ``` sentieon driver -t NUMBER_THREADS -r REFERENCE \ -i TUMOR_DEDUPED_BAM [-q TUMOR_RECAL_DATA.TABLE] \ --algo TNscope --tumor_sample TUMOR_SAMPLE_NAME \ --pon PANEL_OF_NORMAL_VCF [--dbsnp DBSNP] OUT_TN_VCF ``` 该命令需要以下输入: * 线程数:计算中使用的计算机线程数。建议该数值不要超过系统可用计算核心数。 * 参考文件:参考 FASTA 文件的位置。请确保此参考文件与映射阶段使用的文件相同。 * `TUMOR_DEDUPED_BAM`:肿瘤样本去重后预处理 BAM 文件的位置。 * `NORMAL_DEDUPED_BAM`:正常样本去重后预处理 BAM 文件的位置。 * `TUMOR_SAMPLE_NAME` :用于将 reads 比对到参考阶段的肿瘤样本的样本名称。 * `OUT_TN_VCF` :包含变异体的输出文件的位置和文件名。 以下输入参数为可选: * `TUMOR_RECAL_DATA.TABLE`:肿瘤样本的 BQSR 阶段结果存储位置。 * `NORMAL_RECAL_DATA.TABLE`:Normal 样本的 BQSR 阶段结果存储位置。 * `NORMAL_SAMPLE_NAME` :在将读取数据映射到参考序列阶段使用的正常样本名称。 * `DBSNP`:单核苷酸多态性数据库(dbSNP)的位置。dbSNP 中的变异更有可能被标记为种系变异,因为它们需要更多证据证明在正常人群中不存在。您只能使用一个 dbSNP 文件。 * `PANEL_OF_NORMAL_VCF` :Panel of Normal VCF 文件的位置和名称。 ## 4. 生成 PON VCF 为了生成用于 TNscope® 的 PON VCF 文件,需要对 panel 中要使用的每个正常样本运行以下命令: ``` sentieon driver -t NUMBER_THREADS -r REFERENCE -i NORMAL_DEDUPED_BAM \ [-q NORMAL_RECAL_DATA.TABLE] --algo TNscope \ --tumor_sample NORMAL_SAMPLE_NAME OUT_NORMAL_VCF ``` 该命令需要以下输入: * 线程数:计算中使用的计算机线程数。建议该数值不要超过系统可用计算核心数。 * 参考文件:参考 FASTA 文件的位置。请确保此参考文件与映射阶段使用的文件相同。 * `NORMAL_DEDUPED_BAM` :正常样本去重后预处理 BAM 文件的位置。 * `NORMAL_SAMPLE_NAME`:在将读取数据映射到参考序列阶段使用的正常样本名称。 * `OUT_NORMAL_VCF` :包含输入正常样本相关变异的输出 VCF 文件的位置和名称。 以下输入参数为可选: - `NORMAL_RECAL_DATA.TABLE` :Normal 样本的 BQSR 阶段结果存储位置。 生成所有要包含在面板中的 VCF 文件后,需要将它们合并成一个单独的 Normal VCF 面板。可以使用 bcftools 完成此操作: ``` BCF=/path_to_bcftools export BCFTOOLS_PLUGINS=$BCF/plugins DIR=/path_to_normal_vcf_file $BCF/bcftools merge -m all -f PASS,. --force-samples $DIR/*.vcf.gz |$BCF/bcftools plugin fill-AN-AC |$BCF/bcftools filter -i 'SUM(AC)>1' > panel_of_normal.vcf ``` **官方文档**: <https://support.sentieon.com/docs/TNscope_usage/tnscope/#> **Sentieon TNscope 流程 GitHub 地址**: <https://github.com/Insvast/bioinformatics/tree/main/TNscope_WGS> *** # 三、实际测试运行 本次测序运行使用 GIAB 项目的 HG008 样本和真实生产项目中的 Test 样本的 FASTQ 数据作为输入,从而进行 TNscope 分析查看实际的运行效率。 ## 1. Sentieon 软件下载安装 - X86 架构 CPU 服务器版本,例如 Intel、AMD、曙光:https://ftp.insvast.com/user/Sentieon/release/sentieon-genomics-202503.03.tar.gz - ARM 架构 CPU 服务器版本,例如华为鲲鹏、阿里倚天、Ampere:https://ftp.insvast.com/user/Sentieon/release/arm-sentieon-genomics-202503.03.tar.gz 软件下载链接需用户名与密码,用户名:insvast;密码:Ins@1234 ## 2. 测试项目使用配置 - CPU:Intel(R) Xeon(R) Platinum 8358P CPU @ 2.60GHz,64 核心 - 内存:512GB DDR4 - 系统:Ubuntu 22.04.3 LTS ## 3. 参考基因组和 dbSNP 下载 ``` curl -L \ -o 'hg38_ucsc.fa' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/WholeGenomeFasta/genome.fa' \ -o 'hg38_ucsc.fa.fai' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/WholeGenomeFasta/genome.fa.fai' \ -o 'hg38_ucsc.fa.amb' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/BWAIndex/genome.fa.amb' \ -o 'hg38_ucsc.fa.ann' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/BWAIndex/genome.fa.ann' \ -o 'hg38_ucsc.fa.bwt' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/BWAIndex/genome.fa.bwt' \ -o 'hg38_ucsc.fa.pac' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/BWAIndex/genome.fa.pac' \ -o 'hg38_ucsc.fa.sa' 'https://ngi-igenomes.s3.amazonaws.com/igenomes/Homo_sapiens/UCSC/hg38/Sequence/BWAIndex/genome.fa.sa' \ -O 'https://storage.googleapis.com/gcp-public-data--broad-references/hg38/v0/Homo_sapiens_assembly38.dbsnp138.vcf.gz' \ -O 'https://storage.googleapis.com/gcp-public-data--broad-references/hg38/v0/Homo_sapiens_assembly38.dbsnp138.vcf.gz.tbi' ``` ## 4. 测试数据下载 ``` # HG008 来自 GIAB 项目 wget -c https://42basepairs.com/download/web/giab/data_somatic/HG008/Liss_lab/superseded-2022-data/BCM_ILMN-somatic-analysis_20220816/HG008-N_Illumina_R1.fastq.gz wget -c https://42basepairs.com/download/web/giab/data_somatic/HG008/Liss_lab/superseded-2022-data/BCM_ILMN-somatic-analysis_20220816/HG008-N_Illumina_R2.fastq.gz wget -c https://42basepairs.com/download/web/giab/data_somatic/HG008/Liss_lab/superseded-2022-data/BCM_ILMN-somatic-analysis_20220816/HG008-T_Illumina_R1.fastq.gz wget -c https://42basepairs.com/download/web/giab/data_somatic/HG008/Liss_lab/superseded-2022-data/BCM_ILMN-somatic-analysis_20220816/HG008-T_Illumina_R2.fastq.gz # Test 样本为私有样本,不提供下载 ``` ## 5. 分析运行 ``` sbatch somatic_wgs_TNscope.HG008.sh sbatch somatic_wgs_TNscope.Test.sh ``` ## 6. 分析结果展示 ``` ├── [ 10M] output_tnscope.filtered.vcf.gz ├── [146K] output_tnscope.filtered.vcf.gz.tbi ├── [ 10M] output_tnscope.pre_filter.vcf.gz ├── [146K] output_tnscope.pre_filter.vcf.gz.tbi ├── [ 32K] run.log ├── [1.2K] tumor_aln_metrics.txt ├── [ 79G] tumor_coverage_metrics ├── [8.5K] tumor_coverage_metrics.sample_cumulative_coverage_counts ├── [6.3K] tumor_coverage_metrics.sample_cumulative_coverage_proportions ├── [6.9K] tumor_coverage_metrics.sample_interval_statistics ├── [ 14K] tumor_coverage_metrics.sample_interval_summary ├── [ 12K] tumor_coverage_metrics.sample_statistics ├── [ 160] tumor_coverage_metrics.sample_summary ├── [135G] tumor_deduped.bam ├── [9.0M] tumor_deduped.bam.bai ├── [1.9K] tumor_dedup_metrics.txt ├── [6.0K] tumor_gc_metrics.txt ├── [ 534] tumor_gc_summary.txt ├── [ 12K] tumor_is_metrics.txt ├── [4.3K] tumor_mq_metrics.txt ├── [ 295] tumor_qd_metrics.txt ├── [2.8G] tumor_score.txt ├── [3.0M] tumor_score.txt.idx ├── [134G] tumor_sorted.bam ├── [9.0M] tumor_sorted.bam.bai ├── [3.8K] tumor.wgs_metrics.txt ``` ## 7. 质控统计 | 指标 | Test-normal | Test-tumor | HG008-normal | HG008-tumor | |---|---|---|---|---| | TotalReads | 890057280 | 770464744 | 3124709250 | 2912610202 | | TotalBases(Gb) | 133.508592 | 115.569712 | 471.831097 | 439.804141 | | MappedRate(%) | 99.9773 | 99.9553 | 99.6472 | 99.7546 | | Duplicate(%) | 0.8671 | 1.7955 | 20.6579 | 20.1688 | | MeanDepth | 27.563 | 21.426 | 142.41 | 133.85 | | PCT_1X | 97.258 | 96.595 | 97.472 | 97.242 | | PCT_5X | 96.208 | 95.013 | 97.056 | 96.842 | | PCT_10X | 94.35 | 89.959 | 96.861 | 96.649 | | PCT_15X | 89.95 | 78.538 | 96.723 | 96.511 | | PCT_20X | 82.096 | 59.534 | 96.606 | 96.391 | | PCT_25X | 68.038 | 36.153 | 96.502 | 96.281 | | PCT_30X | 45.819 | 16.976 | 96.405 | 96.176 | | PCT_40X | 8.0265 | 2.5083 | 96.218 | 95.962 | | PCT_50X | 0.58713 | 0.72416 | 96.029 | 95.689 | | PCT_60X | 0.21731 | 0.36173 | 95.819 | 95.155 | | PCT_70X | 0.15295 | 0.23001 | 95.56 | 93.4 | | PCT_80X | 0.11544 | 0.15452 | 95.237 | 87.976 | | PCT_90X | 0.092602 | 0.10481 | 94.813 | 77.468 | | PCT_100X | 0.076159 | 0.073361 | 94.122 | 65.48 | | MedianInsertSize | 190 | 165 | 438 | 420 | ## 8. 耗时统计 | 步骤 | Test | HG008 | |---|---|---| | Normal 比对+排序时间(min) | 37.49 | 379.43 | | Normal 去重时间(min) | 4.68 | 10.37 | | Normal 统计 metrics 质控信息时间(min) | 2.41 | 6.12 | | Normal LocusCollector 统计 | 1.13 | 3.67 | | Tumor 比对+排序时间(min) | 71.53 | 406.08 | | Tumor 去重时间(min) | 5.67 | 11.38 | | Tumor 统计 metrics 质控信息时间(min) | 2.65 | 6.57 | | Tumor LocusCollector 统计 | 1.51 | 4.72 | | TNscope 变异检测时间(min) | 30.69 | 9.02 | | 总时间 | 157.75 | 837.35 | **在 64 核测试服务器上,tumor 115Gb+normal 133Gb 数据量的人类 WGS 数据(27.5X Tumor + 21.4X Normal WGS 样本)最快分析仅耗时 157.75 分钟。若 TNscope 前面步骤同时跑,时间能进一步压缩到 112 分钟,极大缩短了分析时间,加快科研成果转化。** Sentieon 在不断地优化算法的运行效率,为科研工作者提供更快速、更经济的基因检测方案。若您刚好有需要检测的数据,不妨来申请试用 Sentieon。 *** # Sentieon 软件介绍 Sentieon 为完整的纯软件基因变异检测二级分析方案,其分析流程完全忠于 BWA、GATK、MuTect2、STAR、Minimap2、Fgbio、picard 等金标准的数学模型。在匹配开源流程分析结果的前提下,大幅提升 WGS、WES、Panel、UMI、ctDNA、RNA 等测序数据的分析效率和检出精度,并匹配目前全部第二代、三代测序平台。 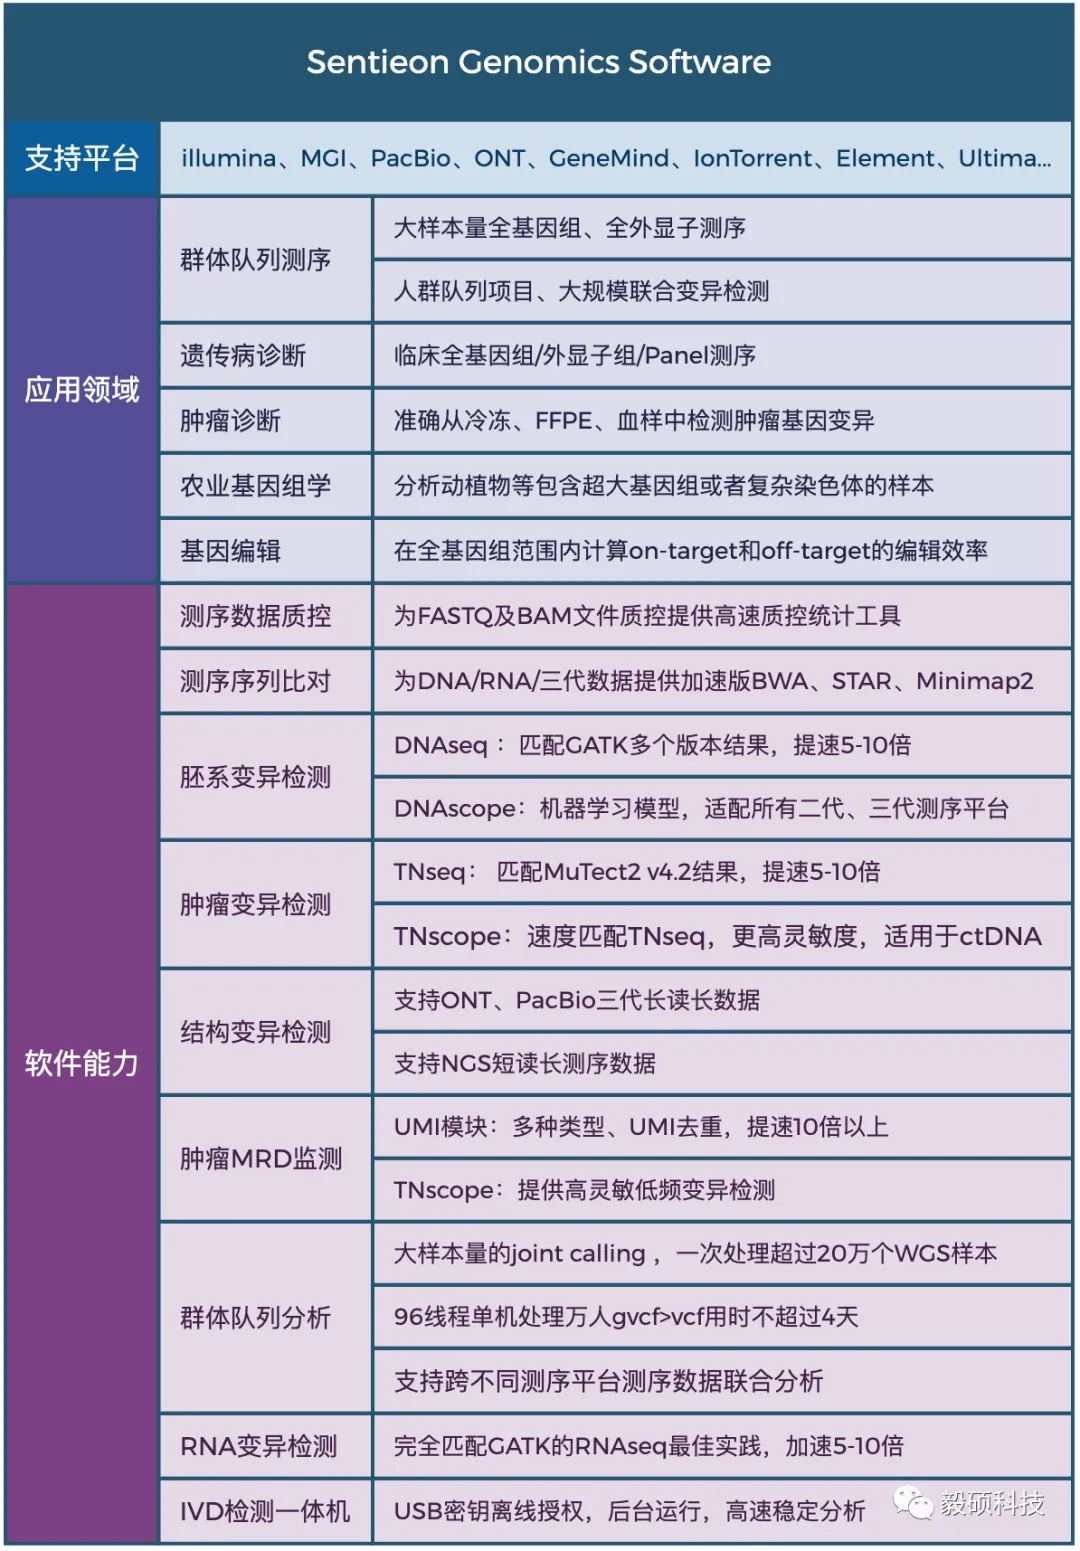 Sentieon 软件团队拥有丰富的软件开发及算法优化工程经验,致力于解决生物数据分析中的速度与准确度瓶颈,**为来自于分子诊断、药物研发、临床医疗、人群队列、动植物等多个领域的合作伙伴提供高效精准的软件解决方案,共同推动基因技术的发展。**** 截至 2026 年 4 月份,Sentieon 已经在全球范围内为 1860+ 用户提供服务,用户处理超过 7400+ PB 数据量,被世界一级影响因子刊物如 NEJM、Cell、Nature 等广泛引用,引用次数超过 1900 篇。此外,Sentieon 连续数年摘得了 Precision FDA、Dream Challenges 等多个权威评比的桂冠,在业内获得广泛认可。
chsnp
May 27, 2026, 5:29 p.m.
Forward
Favorites
Last
Next
Scan the QR Code
Copy link
Scan the QR code to share.
Copy link
Markdown file
Word document
PDF document
PDF document (print)
share
link
type
password
Update password
Validity period